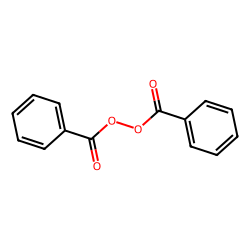
Cheméo RefData 1.0
One consistent value set for simulation, built from chemeo v31.
- Formula
- C14H10O4
- Molecular Weight
- 242.23 g/mol
- SMILES
- O=C(OOC(=O)c1ccccc1)c1ccccc1
- InChI Key
- OMPJBNCRMGITSC-UHFFFAOYSA-N
Cheméo is a service of Céondo GmbH, since 2007, we provide simulation, modelling and software development services for the industry.
Do not hesitate to contact us for your projects.
Constants
| Property | Value | Unit | u95 | Basis | Sources |
|---|---|---|---|---|---|
| MW Molecular Weight |
242.23 | g/mol | derived | ||
| Tc Critical Temperature |
884.00 | K | compiled Compiled value | ChemSep v8.3 pure component data | |
| Pc Critical Pressure |
2570.00 | kPa | compiled Compiled value | ChemSep v8.3 pure component data | |
| Vc Critical Volume |
0.643 | m3/kmol | compiled Compiled value | ChemSep v8.3 pure component data | |
| Zc Critical Compressibility |
0.2248318 | derived | |||
| ω Acentric Factor |
0.9136 | derived | |||
| Tboil Normal Boiling Point Temperature |
669.46 | K | derived | ||
| Tfus Normal melting (fusion) point |
380.00 | K | compiled Compiled value | ChemSep v8.3 pure component data | |
| ΔfG° Standard Gibbs free energy of formation |
-108.80 | kJ/mol | compiled Compiled value | ChemSep v8.3 pure component data |
Equations
| Property | Equation | Temperature (K) | Fit |
|---|---|---|---|
| Pvap Vapor pressure |
Y = exp(A + B/T + C·ln(T) + D·T^E) Y in kPa, T in K |
[404.63; 884.00] | compiled, 18 points, AARD 0.0% against its inputs, published equations only: agreement with those equations, not with measurements ChemSep v8.3 pure component data |
| A = 1.13357e+02, B = -1.74600e+04, C = -1.27196e+01, D = 1.13563e-18, E = 6.00000e+00 | |||
| Cp,gas Ideal gas heat capacity |
Y = A + B·((C/T)/sinh(C/T))² + D·((E/T)/cosh(E/T))² Y in J/mol×K, T in K |
[255.00; 1500.00] | compiled, 20 points, AARD 0.2% against its inputs, published equations only: agreement with those equations, not with measurements ChemSep v8.3 pure component data |
| A = 9.92424e+01, B = 5.65098e+02, C = 1.55109e+03, D = 4.11498e+02, E = 6.90711e+02 | |||
| Cp,liquid Liquid phase heat capacity |
Y = A + B·T + C·T² + D·T³ + E·T⁴ Y in J/mol×K, T in K |
[442.00; 662.00] | compiled, 20 points, AARD 0.0% against its inputs, published equations only: agreement with those equations, not with measurements ChemSep v8.3 pure component data |
| A = 8.21000e+01, B = 6.42200e-01, C = -2.31555e-16, D = 1.34947e-19, E = 0.00000e+00 | |||
| ΔvapH Enthalpy of vaporization at a given temperature |
Y = A·(1 − Tr)^(B + C·Tr + D·Tr² + E·Tr³), Tr = T/Tc Y in kJ/mol, T in K |
[404.63; 857.37] | derived, 27 points, AARD 0.2% against its inputs ChemSep v8.3 pure component data |
| A = 1.33514e+02, B = 5.13072e-01, C = -1.68565e-01, D = 5.27619e-02, E = 0.00000e+00, Tc = 8.84000e+02 K | |||
| ρl Liquid Density |
Y = A / B^(1 + (1 − T/C)^D) Y in kg/m3, T in K |
[404.63; 857.37] | compiled, 18 points, AARD 0.0% against its inputs, published equations only: agreement with those equations, not with measurements ChemSep v8.3 pure component data |
| A = 8.48285e+01, B = 2.51099e-01, C = 8.84000e+02, D = 2.86000e-01 | |||
| λliquid Liquid thermal conductivity |
Y = A + B·T + C·T² + D·T³ + E·T⁴ Y in W/m×K, T in K |
[393.32; 669.00] | compiled, 19 points, AARD 0.0% against its inputs, published equations only: agreement with those equations, not with measurements ChemSep v8.3 pure component data |
| A = 1.47000e-01, B = -1.22320e-04, C = 4.27535e-20, D = -2.55092e-23, E = 0.00000e+00 | |||
| λgas Gas thermal conductivity |
Y = A·T^B / (1 + C/T + D/T²) Y in W/m×K, T in K |
[669.00; 1000.00] | compiled, 20 points, AARD 0.1% against its inputs, published equations only: agreement with those equations, not with measurements ChemSep v8.3 pure component data |
| A = 1.51643e-06, B = 1.47438e+00, C = 0.00000e+00, D = 0.00000e+00 | |||
| γ Surface Tension |
Y = A·(1 − Tr)^(B + C·Tr + D·Tr² + E·Tr³), Tr = T/Tc Y in N/m, T in K |
[404.63; 857.37] | compiled, 18 points, AARD 0.0% against its inputs, published equations only: agreement with those equations, not with measurements ChemSep v8.3 pure component data |
| A = 6.69200e-02, B = 1.20330e+00, C = -6.08084e-14, D = 1.11702e-14, E = 0.00000e+00, Tc = 8.84000e+02 K | |||
Equation of state
| Model | Tc (K) | Pc (kPa) | ω | c1 | c2 | c3 | Volume shift (m³/kmol) | Pvap AARD |
|---|---|---|---|---|---|---|---|---|
| PR | 884.00 | 2570.00 | 0.9136 | 1.66967e+00 | -4.48630e-01 | 6.80672e-01 | 1.62095e-03 | 0.4% |
| SRK | 884.00 | 2570.00 | 0.9136 | 1.90225e+00 | -9.50995e-01 | 1.30708e+00 | 3.66849e-02 | 0.6% |
a(T) = Ωa·R²·Tc²/Pc·α(T) and b = Ωb·R·Tc/Pc, with Pc in Pa and R = 8.314462618 J/(mol·K); PR: Ωa = 0.45724, Ωb = 0.07780; SRK: Ωa = 0.42748, Ωb = 0.08664.
Mathias-Copeman alpha function, with Tr = T/Tc:
α(T) = [1 + c1·(1 − √Tr) + c2·(1 − √Tr)² + c3·(1 − √Tr)³]² for T ≤ Tc,
α(T) = [1 + c1·(1 − √Tr)]² above Tc.
Péneloux volume shift: v = vEOS − volume shift, fitted to the liquid density at 0.7·Tc (0 when there is no liquid density).
A binary interaction parameter is valid only with these parameters, from RefData 1.0.
Not in RefData for this compound
Enthalpy of formation at standard conditions, Rackett Parameter, Dynamic viscosity.