Cheméo RefData 1.0
One consistent value set for simulation, built from chemeo v31.
- Formula
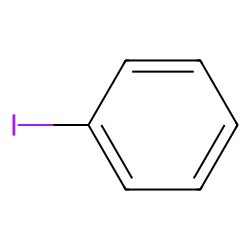
- C6H5I
- CAS
- 591-50-4
- Molecular Weight
- 204.01 g/mol
- SMILES
- Ic1ccccc1
- InChI Key
- SNHMUERNLJLMHN-UHFFFAOYSA-N
Constants
| Property | Value | Unit | u95 | Basis | Sources |
|---|---|---|---|---|---|
| MW Molecular Weight |
204.01 | g/mol | derived | ||
| Tc Critical Temperature |
721.00 | K | 8.65 | experimental | KDB |
| Pc Critical Pressure |
4520.00 | kPa | 244.08 | experimental | KDB |
| Vc Critical Volume |
0.351 | m3/kmol | 0.021 | experimental | KDB |
| Zc Critical Compressibility |
0.2646526 | derived | |||
| ω Acentric Factor |
0.2465 | derived | |||
| Tboil Normal Boiling Point Temperature |
461.60 | K | 0.54 | experimental | KDB; NIST Webbook |
| Tfus Normal melting (fusion) point |
241.90 | K | 0.46 | experimental | KDB; Aqueous Solubility Prediction Method; NIST Webbook |
| Zra Rackett Parameter |
0.26 | 0.01 | experimental | KDB |
Equations
| Property | Equation | Temperature (K) | Fit |
|---|---|---|---|
| Pvap Vapor pressure |
Y = exp(A + B/T + C·ln(T) + D·T^E) Y in kPa, T in K |
[241.83; 721.00] | experimental, 67 points, AARD 0.9% against the measurements |
| A = 1.00908e+02, B = -8.98676e+03, C = -1.33145e+01, D = 1.05282e-02, E = 1.00000e+00 | |||
| Cp,gas Ideal gas heat capacity |
Y = A + B·((C/T)/sinh(C/T))² + D·((E/T)/cosh(E/T))² Y in J/mol×K, T in K |
[200.00; 1500.00] | compiled, 20 points, AARD 0.2% against its inputs, published equations only: agreement with those equations, not with measurements ChemSep v8.3 pure component data |
| A = 6.81200e+01, B = 2.04718e+02, C = 1.39810e+03, D = 1.42211e+02, E = 6.66159e+02 | |||
| Cp,liquid Liquid phase heat capacity |
Y = A + B·T + C·T² + D·T³ + E·T⁴ Y in J/mol×K, T in K |
[250.00; 320.00] | experimental, 12 points, AARD 0.4% against the measurements |
| A = -4.54350e+02, B = 6.36254e+00, C = -2.24230e-02, D = 2.68162e-05, E = 0.00000e+00 | |||
| ΔvapH Enthalpy of vaporization at a given temperature |
Y = A·(1 − Tr)^(B + C·Tr + D·Tr² + E·Tr³), Tr = T/Tc Y in kJ/mol, T in K |
[241.83; 695.92] | derived, 29 points, AARD 0.6% against its inputs ChemSep v8.3 pure component data |
| A = 5.88128e+01, B = 2.55081e-01, C = 2.75899e-01, D = -1.62439e-01, E = 0.00000e+00, Tc = 7.21000e+02 K | |||
| ρl Liquid Density |
Y = A / B^(1 + (1 − T/C)^D) Y in kg/m3, T in K |
[241.83; 695.92] | compiled, 20 points, AARD 0.0% against its inputs, published equations only: agreement with those equations, not with measurements ChemSep v8.3 pure component data |
| A = 1.54889e+02, B = 2.64985e-01, C = 7.21000e+02, D = 2.91000e-01 | |||
| η Dynamic viscosity |
Y = exp(A + B/T + C·ln(T) + D·T^E) Y in Pa×s, T in K |
[241.83; 461.60] | compiled, 20 points, AARD 0.8% against its inputs, published equations only: agreement with those equations, not with measurements ChemSep v8.3 pure component data |
| A = -2.59041e+01, B = 2.06836e+03, C = 2.19673e+00, D = 0.00000e+00, E = 0.00000e+00 | |||
| λliquid Liquid thermal conductivity |
Y = A + B·T + C·T² + D·T³ + E·T⁴ Y in W/m×K, T in K |
[242.17; 461.60] | compiled, 18 points, AARD 0.1% against its inputs, published equations only: agreement with those equations, not with measurements ChemSep v8.3 pure component data |
| A = 8.18790e-02, B = 3.48136e-04, C = -1.32219e-06, D = 1.23069e-09, E = 0.00000e+00 | |||
| λgas Gas thermal conductivity |
Y = A·T^B / (1 + C/T + D/T²) Y in W/m×K, T in K |
[241.83; 1000.00] | compiled, 20 points, AARD 2.8% against its inputs, published equations only: agreement with those equations, not with measurements ChemSep v8.3 pure component data |
| A = 1.39369e-05, B = 1.21063e+00, C = 4.14205e+02, D = 5.32478e+03 | |||
| γ Surface Tension |
Y = A·(1 − Tr)^(B + C·Tr + D·Tr² + E·Tr³), Tr = T/Tc Y in N/m, T in K |
[241.83; 666.84] | compiled, 39 points, AARD 0.5% against its inputs, published equations only: agreement with those equations, not with measurements Jasper (1972), surface tension of pure liquid compounds; ChemSep v8.3 pure component data |
| A = 6.87807e-02, B = 7.34273e-01, C = 1.07977e+00, D = -6.26881e-01, E = 0.00000e+00, Tc = 7.21000e+02 K | |||
Equation of state
| Model | Tc (K) | Pc (kPa) | ω | c1 | c2 | c3 | Volume shift (m³/kmol) | Pvap AARD |
|---|---|---|---|---|---|---|---|---|
| PR | 721.00 | 4520.00 | 0.2465 | 7.08814e-01 | 1.63932e-01 | 1.39327e-02 | -6.71882e-04 | 0.0% |
| SRK | 721.00 | 4520.00 | 0.2465 | 8.69088e-01 | -1.27400e-01 | 3.39064e-01 | 1.73104e-02 | 0.0% |
a(T) = Ωa·R²·Tc²/Pc·α(T) and b = Ωb·R·Tc/Pc, with Pc in Pa and R = 8.314462618 J/(mol·K); PR: Ωa = 0.45724, Ωb = 0.07780; SRK: Ωa = 0.42748, Ωb = 0.08664.
Mathias-Copeman alpha function, with Tr = T/Tc:
α(T) = [1 + c1·(1 − √Tr) + c2·(1 − √Tr)² + c3·(1 − √Tr)³]² for T ≤ Tc,
α(T) = [1 + c1·(1 − √Tr)]² above Tc.
Péneloux volume shift: v = vEOS − volume shift, fitted to the liquid density at 0.7·Tc (0 when there is no liquid density).
A binary interaction parameter is valid only with these parameters, from RefData 1.0.
Not in RefData for this compound
Enthalpy of formation at standard conditions, Standard Gibbs free energy of formation.
Checks
Deviation from the compiled data (ChemSep, ChEDL): Critical Temperature: 0.0%; Critical Pressure: 0.0%; Critical Volume: 0.0%; Normal Boiling Point Temperature: 0.0%; Normal melting (fusion) point: 0.0%; Vapor pressure: 0.6%; Liquid phase heat capacity: 1.9%.